

Get minimum distances#
Calculating minimum spatial distances between atom selections or molecular groups.
The function molsysmt.structure.get_minimum_distances() calculates the closest distance and identifies the closest atom pair between two groups of atoms across single or multiple trajectory structures.
Added in version 1.0.0.
API documentation
Follow this link for a detailed description of the input arguments, raised errors, and returned objects of this function: molsysmt.structure.get_minimum_distances().
Basic usage#
Let’s show how to calculate the minimum distance between two atom sets over a pentalanine trajectory (illustrated using terminal residues 0 and 4):
import molsysmt as msm
import matplotlib.pyplot as plt
import pyunitwizard as puw
molsys = msm.convert(msm.systems['pentalanine']['traj_pentalanine.h5msm'])
Note
Demo Systems Catalog
This tutorial uses demonstration datasets provided by MolSysMT. To explore the full catalog of bundled systems, forms, and file paths, visit the Demo Systems guide.
We compute the minimum distance between the two selections using molsysmt.structure.get_minimum_distances():
pairs, min_distances = msm.structure.get_minimum_distances(molsys, selection='group_index==0', selection_2='group_index==4')
print('Pairs array shape:', pairs.shape)
print('Minimum distances shape:', min_distances.shape)
print(f'Minimum distance at frame 0: {min_distances[0]:.3f} (between atom {pairs[0,0]} and {pairs[0,1]})')
Pairs array shape: (5000, 2)
Minimum distances shape: (5000,)
Minimum distance at frame 0: 0.823 nanometer (between atom 4 and 3)
Plotting minimum distance time series#
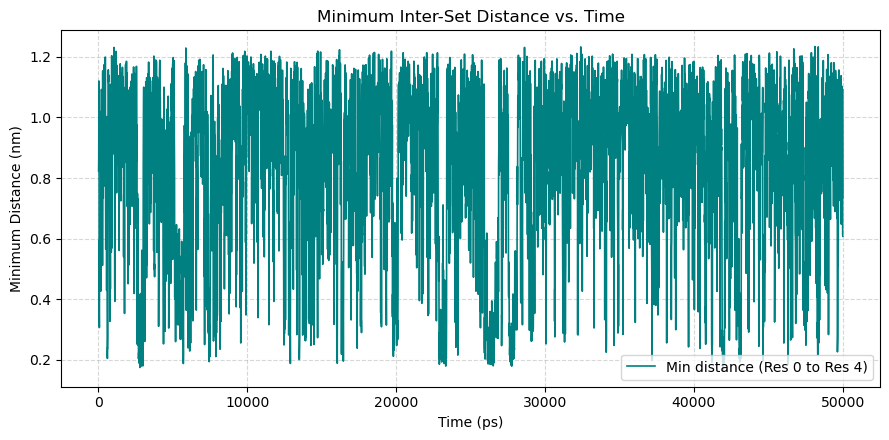
We plot the evolution of the minimum inter-set distance over simulation time:
time = msm.get(molsys, element='system', time=True)
time_val = puw.get_value(time, to_unit='ps')
dist_val = puw.get_value(min_distances, to_unit='nm')

plt.figure(figsize=(9, 4.5))
plt.plot(time_val, dist_val, color='teal', lw=1.2, label='Min distance (Res 0 to Res 4)')
plt.xlabel('Time (ps)')
plt.ylabel('Minimum Distance (nm)')
plt.title('Minimum Inter-Set Distance vs. Time')
plt.grid(True, linestyle='--', alpha=0.5)
plt.legend()
plt.tight_layout()
plt.show()
Finding the closest atom in a target set#
Setting as_entity=False and as_entity_2=True queries, for each atom in selection, the closest atom in selection_2:
pairs_ind, min_d_ind = msm.structure.get_minimum_distances(molsys, selection='group_index==0 and atom_type!="H"', selection_2='group_index==4 and atom_type!="H"', as_entity=False, as_entity_2=True)
print('Shape of individual atom minimum distances:', min_d_ind.shape)
print(f'Closest target atom for first atom at frame 0: target atom index {pairs_ind[0, 0]} at distance {min_d_ind[0, 0]:.3f}')
Shape of individual atom minimum distances: (5000, 3)
Closest target atom for first atom at frame 0: target atom index 0 at distance 0.977 nanometer
Computing minimum distances between atom sets#
We can compute the global minimum non-bonded distance between two independent macro-selections (illustrated here using two protein chains in TcTIM):
tctim = msm.convert(msm.systems['TcTIM']['1tcd.h5msm'])
pairs_chains, min_dist_chains = msm.structure.get_minimum_distances(tctim, selection='chain_index==0 and atom_type!="H"', selection_2='chain_index==1 and atom_type!="H"')
print(f'Closest inter-chain contact distance: {min_dist_chains[0]:.3f}')
print(f'Closest contact atoms: atom {pairs_chains[0,0]} (chain 0) and atom {pairs_chains[0,1]} (chain 1)')
Closest inter-chain contact distance: 0.254 nanometer
Closest contact atoms: atom 552 (chain 0) and atom 729 (chain 1)
Computing minimum distances between group centers#
Passing center_of_atoms=True and center_of_atoms_2=True finds the minimum distance between the geometric centroids of atom groups (e.g. between all residue centers of two chains):
groups_0 = msm.get(tctim, element='group', selection='chain_index==0', atom_index=True)
groups_1 = msm.get(tctim, element='group', selection='chain_index==1', atom_index=True)
pairs_cent, min_d_cent = msm.structure.get_minimum_distances(tctim, selection=groups_0, selection_2=groups_1, center_of_atoms=True, center_of_atoms_2=True)
print(f'Closest residue centers distance: {min_d_cent[0]:.3f} (between residue {pairs_cent[0,0]} and {pairs_cent[0,1]})')
Closest residue centers distance: 0.382 nanometer (between residue 69 and 12)
Measuring minimum displacements across structures#
Using pairs=True between different structure indices identifies the atom that experienced the minimum displacement between consecutive time frames:
pairs_disp, min_disp = msm.structure.get_minimum_distances(molsys, structure_indices=[0, 1], structure_indices_2=[1, 2], pairs=True)
print(f'Atom {pairs_disp[0]} had the minimum displacement ({min_disp[0]:.4f}) between frames 0 and 1.')
print(f'Atom {pairs_disp[1]} had the minimum displacement ({min_disp[1]:.4f}) between frames 1 and 2.')
Atom 45 had the minimum displacement (0.0734 nanometer) between frames 0 and 1.
Atom 35 had the minimum displacement (0.2984 nanometer) between frames 1 and 2.
See also
Related Tools & References
Convert: Convert molecular systems between different forms with
molsysmt.basic.convert().Get: Query system time and structural attributes with
molsysmt.basic.get().Get distances: Compute full pairwise distance matrices and element distances with
molsysmt.structure.get_distances().Get maximum distances: Find the maximum span or diameter between elements with
molsysmt.structure.get_maximum_distances().Get contacts: Determine boolean contact matrices within a distance threshold with
molsysmt.structure.get_contacts().